Glycogen storage disease type 1a (GSD1a) is a rare genetic disorder that affects how the body stores and uses carbohydrates for energy.
Under normal circumstances, carbohydrates are consumed through food and converted into two substances, glucose and glycogen.
Glucose is used immediately for energy, while glycogen is stored for later use (mostly in the liver and muscles). Stored glycogen can be converted into glucose to maintain blood sugar levels and provide the body with energy between meals. These are complex biological processes that rely on a variety of different substances interacting with one another at the cellular level.
In people with GSD1a, the body is unable to produce an enzyme called glucose-6-phosphatase (G6Pase). This enzyme plays a critical role in the process where glucose derived from stored glycogen is released into the bloodstream.
As a result, the body cannot use stored glucose. This can lead to dangerously low blood glucose levels (hypoglycemia). Complications can include gout (caused by high uric acid levels), high cholesterol, kidney problems, and liver disease.
The main treatment for GSD1a is cornstarch therapy
Different carbohydrates are digested and metabolized at different rates. At one end of the spectrum, there are simple carbs which are metabolized and used by the body very quickly. Sugar is an example of a simple carb.
At the other end of the spectrum are complex carbs, which are digested and metabolized at a slower rate. One of the slowest digesting/metabolizing carbohydrates is uncooked cornstarch, a common food ingredient derived from corn kernels that is often used as a thickening agent.
Uncooked cornstarch is digested very slowly, which means it acts like a slow-release source of glucose. For someone with GSD1a, frequent feedings that include uncooked cornstarch can help keep blood glucose levels stable.
What type of cornstarch is used?
Cornstarch found in a baking aisle at a grocery store can be used. It is often mixed with cool water or other sugar-free liquids. It cannot be cooked, heated, or mixed with hot liquids, which causes the starch to break down and digest too quickly.
There is also “extended release” cornstarch. These are medical products that have been engineered to digest more slowly than store-bought cornstarch.
Your healthcare provider will give you detailed instructions to follow. Cornstarch therapy requires careful planning, and it can be challenging. Missing or delaying a dose by even a short amount of time can cause a drop in blood glucose. Overnight feedings of cornstarch are needed to maintain blood glucose levels, and a gastric feeding tube may be needed.
What are the other parts of a treatment plan?
Beyond cornstarch therapy, other parts of a GSD1a treatment plan include:
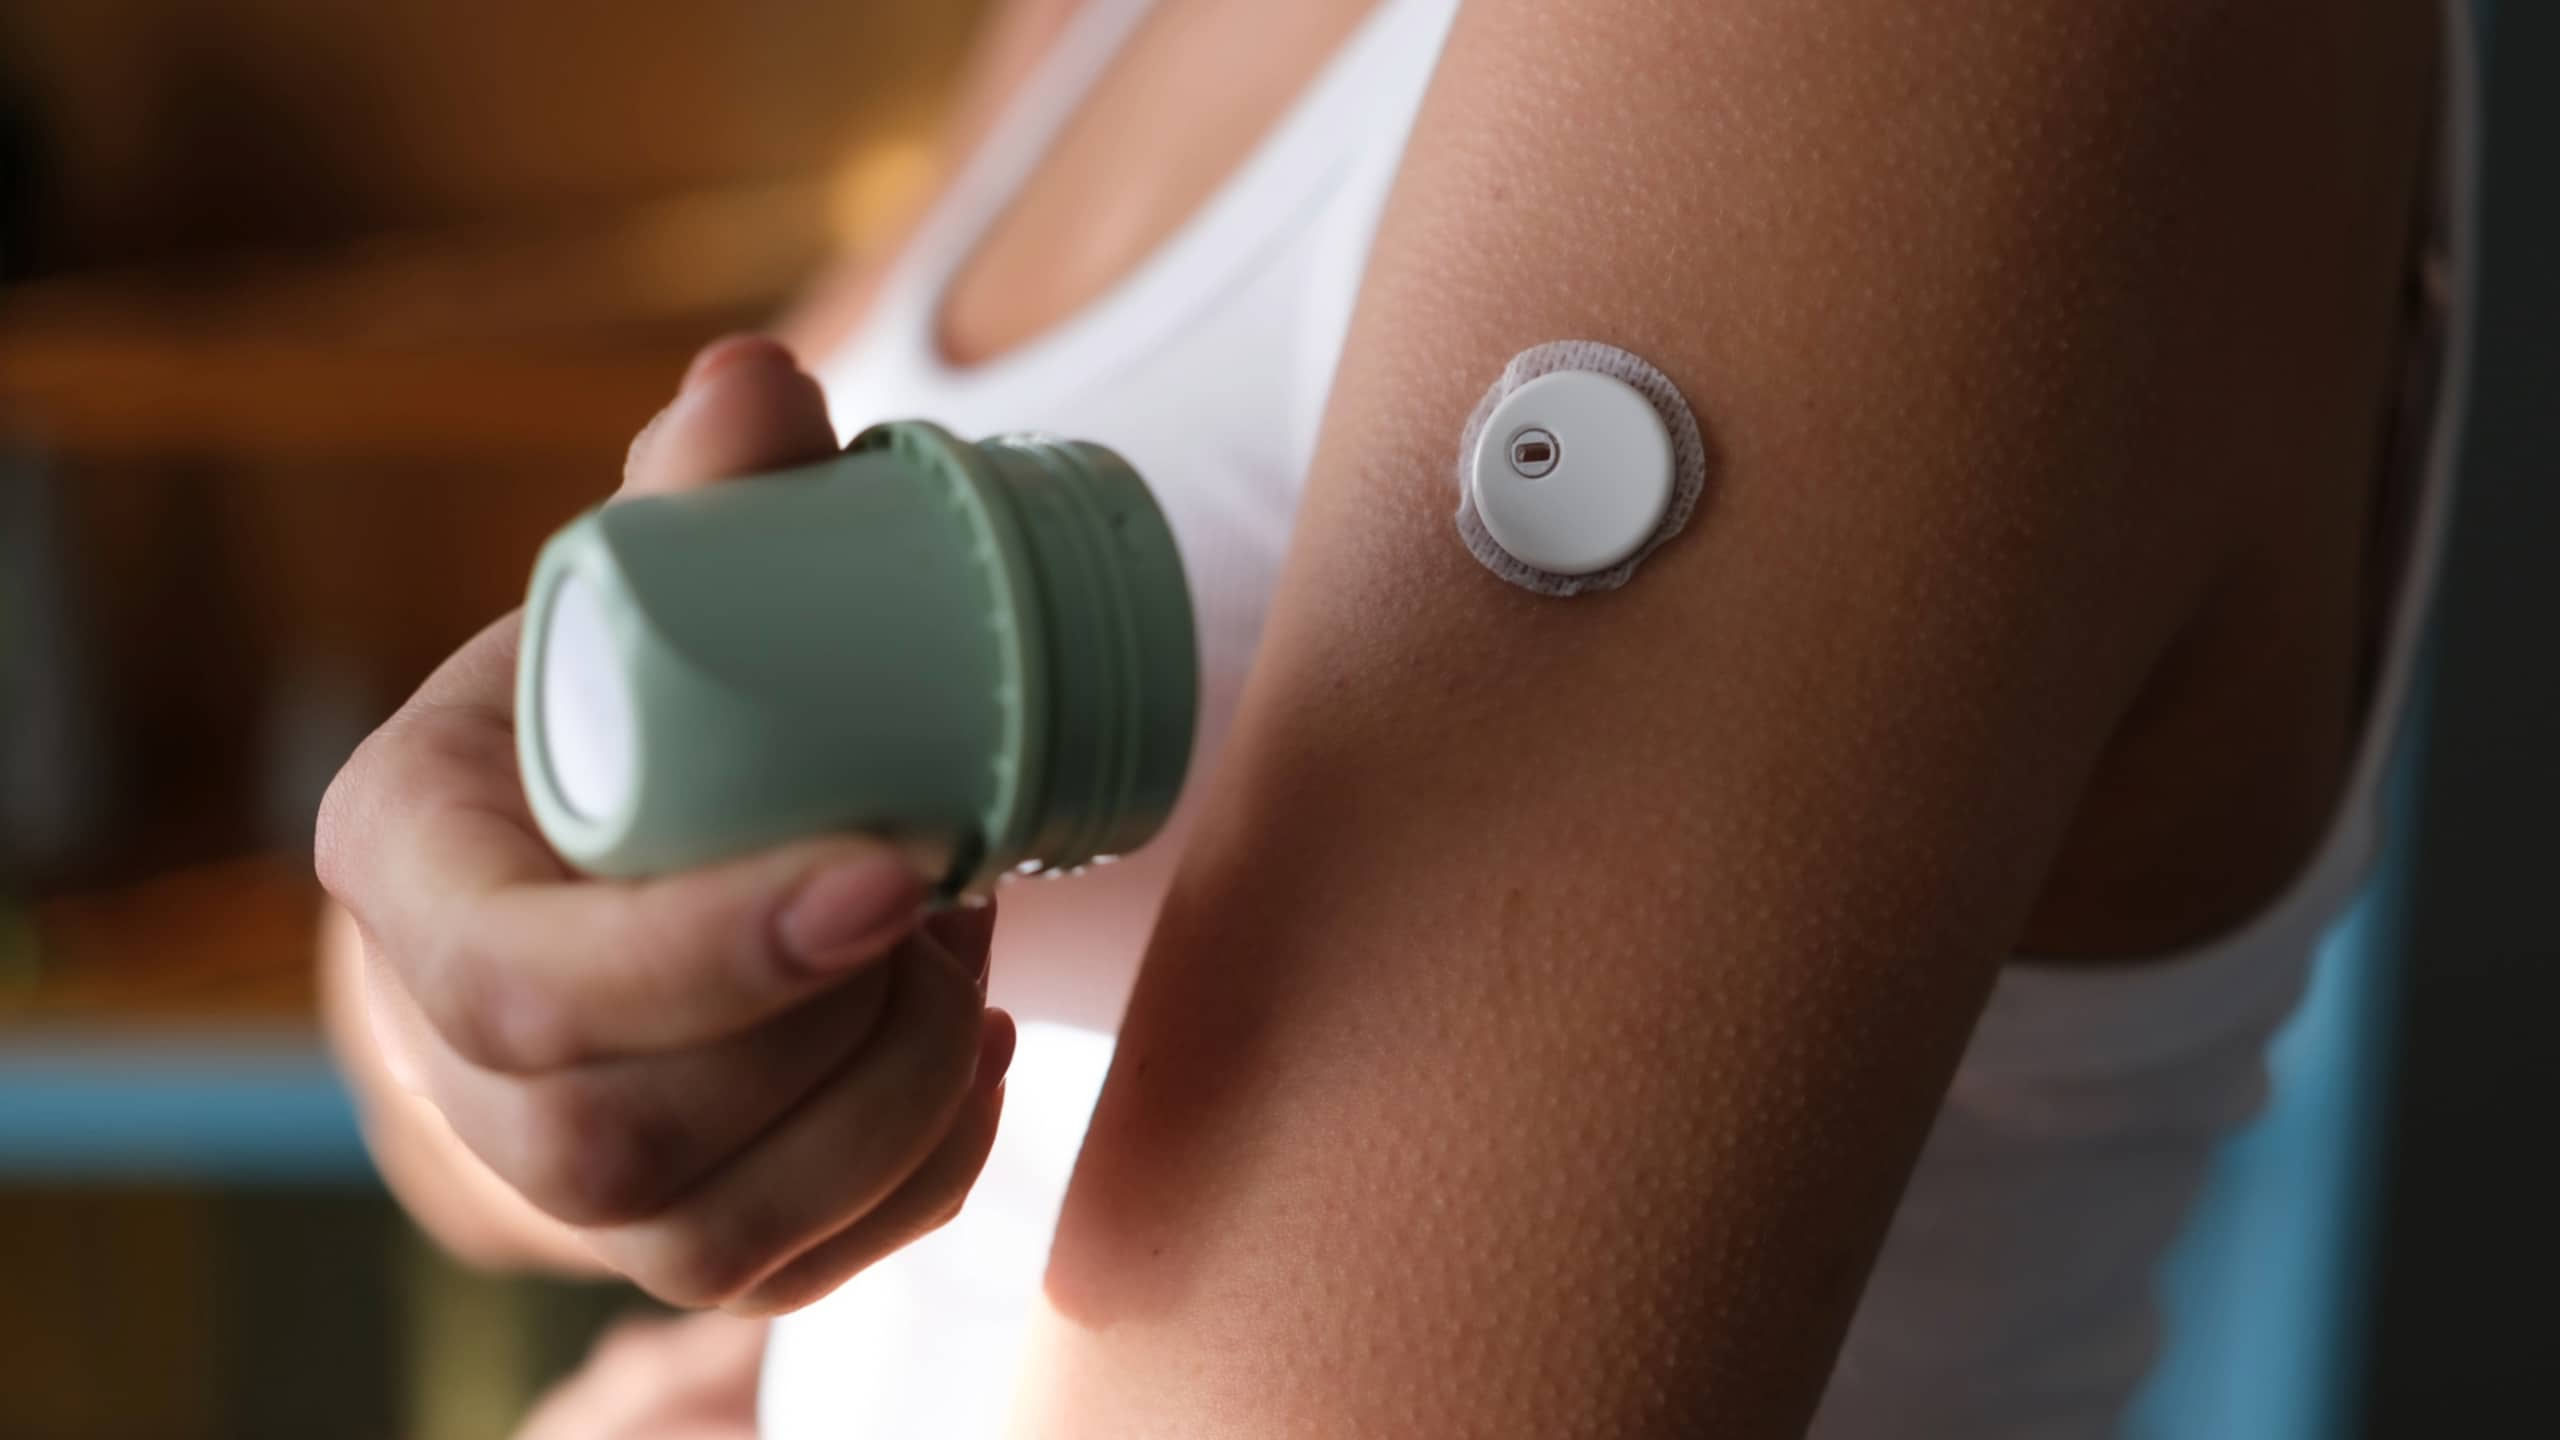
- Using a continuous glucose monitor. This is a small sensor that is attached to the skin to track real-time blood glucose levels.
- Dietary restrictions. Sugars found in dairy products (lactose and galactose), fruit (fructose), and table sugar (sucrose) need to be avoided. GSD1a prevents the body from metabolizing these sugars.
- Additional medications, such as medications to reduce high uric acid levels (to control GSD1a-associated gout), high cholesterol levels, and kidney health.
New and emerging therapies for GSD1a
Medical researchers are actively working on new therapies for glycogen storage disease. Gene therapies are a major focus.
The underlying causes of GSD1a are genetic variations or mutations. These mutations provide faulty instructions for making the G6Pase enzyme the body needs to release glycogen from the cells where it is stored.
A gene therapy provides the body with a functioning version of the faulty gene. If successful, the body will begin to produce the functioning enzyme on its own. This can reduce the need for cornstarch therapy (though it may not eliminate the need completely).
New therapies, emerging therapies, updates to treatment guidelines, and clinical trials are topics worth discussing with your healthcare team, who will be your best source of information.
Sources: National Organization for Rare Disorders... + 12
- National Organization for Rare Disorders. Glycogen Storage Disease Type I. Accessed April 28, 2026.
- Mitan Nana, Catherine Anastasopoulou, et al. Glycogen Storage Disease Type I. StatPearls. April 6, 2025.
- Cleveland Clinic. Glycogen. July 13, 2024.
- Mihir N. Nakrani, Robert H. Wineland, and Fatima Anjum. Physiology, Glucose Metabolism. StatPearls. July 17, 2023.
- Cleveland Clinic. Glycogen Storage Disease (GSD). June 26, 2023.
- American Heart Association. Carbohydrates. September 12, 2023.
- The Children's Fund for Glycogen Storage Disease Research. Daily Management: Diet and cornstarch. Accessed April 28, 2026.
- Terry G. J. Derks, David F. Rodriguez-Buritica, et al. Glycogen Storage Disease Type Ia: Current Management Options, Burden and Unmet Needs. Nutrients, 2021. Vol. 13, No. 11.
- Terry G. J. Derks, Ruben J. Overduin, et al. State of the Art and Consensus Statements by Healthcare Providers, Patients, and Caregivers on Continuous Glucose Monitoring in Liver Glycogen Storage Diseases. Journal of Inherited Metabolic Disorders, 2025. Vol. 48, No. 3.
- The Children's Fund for Glycogen Storage Disease Research. Frequently Asked Questions. Accessed April 28, 2026.
- Kristianne Hannemann. What are new potential treatments in glycogen storage disease type Ia? Drugs.com. March 16, 2026.
- MedlinePlus Genetics. Glycogen storage disease type I. Accessed April 28, 2026.
- Cleveland Clinic. Gene Therapy. July 17, 2025.